


ISO/FDIS 8637-3
(Main)Extracorporeal systems for blood purification — Part 3: Plasmafilters
General Information
- Abstract
This document specifies requirements and test methods for plasmafilters, which are devices intended to separate plasma from blood in therapeutic plasmapheresis therapy. This document specifies the requirements for sterile, single-use plasmafilters, intended for use on humans, hereinafter collectively referred to as “the device”, for use in humans. This document does not apply to; — extracorporeal blood circuits; — haemodialysers, haemodiafilters, haemofilters and haemoconcentrators; — haemoperfusion devices; — vascular access devices; — blood pumps; — systems or equipment intended to perform plasma separation. NOTE 1 Requirements for the extracorporeal blood circuit are specified in ISO 8637-2. NOTE 2 Requirements for haemodialysers, haemodiafilters, haemofilters and haemoconcentrators are specified in ISO 8637-1.
- Status
- Not Published
- Technical Committee
- ISO/TC 150/SC 2 - Cardiovascular implants and extracorporeal systems
- Drafting Committee
- ISO/TC 150/SC 2/WG 5 - Renal replacement, detoxification and apheresis
- Current Stage
- 5000 - FDIS registered for formal approval
- Start Date
- 18-Jun-2026
- Completion Date
- 19-Jan-2026
Buy Documents
ISO/FDIS 8637-3 - Extracorporeal systems for blood purification — Part 3: Plasmafilters
REDLINE ISO/FDIS 8637-3 - Extracorporeal systems for blood purification — Part 3: Plasmafilters
Overview
ISO/FDIS 8637-3: Extracorporeal Systems for Blood Purification - Part 3: Plasmafilters is an international standard developed by the International Organization for Standardization (ISO). This standard specifies requirements and test methods for sterile, single-use plasmafilters designed for use with humans in therapeutic plasmapheresis therapy. Plasmafilters are medical devices intended to separate plasma from blood using a semipermeable membrane, a critical procedure in the management of a range of disorders requiring blood purification.
ISO/FDIS 8637-3 details the essential characteristics of plasmafilters, including biological safety, haemocompatibility, sterility, mechanical strength, and performance criteria. The standard also addresses relevant test methods and labelling requirements, ensuring both safety and consistency across all products intended for human clinical use.
Key Topics
Scope and Exclusions
- Applies to sterile, single-use plasmafilters for human use;
- Does not apply to extracorporeal blood circuits, haemodialysers, haemodiafilters, haemofilters, haemoconcentrators, haemoperfusion devices, vascular access devices, blood pumps, or entire plasma separation systems.
Biological Safety and Haemocompatibility
- Devices must be tested for biocompatibility and absence of biological hazard, referencing ISO 10993 series for biological evaluation of medical devices.
Sterility and Non-pyrogenicity
- All blood and filtrate pathways must be sterile and non-pyrogenic, validated through recognized test methods such as ISO 11737-2 and ISO 10993-11.
Mechanical and Structural Integrity
- Plasmafilters must be able to withstand maximum pressure conditions (positive and negative) encountered during clinical use.
- Connectors must be leak-free and able to resist separation forces and torques as defined by risk management.
Performance Characteristics
- Specifications for plasma filtration rate, sieving coefficients for key solutes (e.g., albumin, immunoglobulins), blood compartment volume, pressure drop, and haemolytic properties.
- Devices must retain safety and performance after storage and up to the stated expiry date.
Labelling and Packaging
- Comprehensive labelling required on the device, unit container, and outer packaging, including instructions for use, identification, and technical details.
Applications
Plasmafilters standardized by ISO/FDIS 8637-3 are essential in therapeutic plasmapheresis, a process where harmful substances are removed from a patient's plasma. This procedure is used in the treatment of:
- Autoimmune disorders
- Neurological diseases
- Hematological conditions
Implementing this standard helps:
- Healthcare providers ensure the safety, reliability, and efficacy of plasmafilters in clinical settings
- Manufacturers demonstrate compliance with internationally recognized quality and safety standards
- Regulatory bodies assess the conformance of new plasmafilters prior to market approval
Adoption of ISO/FDIS 8637-3 facilitates global harmonization, supporting safe patient outcomes and consistent clinical practices worldwide.
Related Standards
ISO/FDIS 8637-3 is part of the broader ISO 8637 series addressing extracorporeal blood purification systems. Complementary and referenced standards include:
- ISO 8637-1: Requirements for haemodialysers, haemodiafilters, haemofilters, and haemoconcentrators
- ISO 8637-2: Requirements for extracorporeal blood circuits
- ISO 14971: Application of risk management to medical devices
- ISO 10993 series: Biological evaluation of medical devices
- ISO 11607-1 / ISO 11607-2: Requirements for packaging of sterilized medical devices
- ISO 11737-2: Sterility testing methods
- ISO 80369-7 / ISO 80369-20: Small-bore connectors in healthcare
Following these related ISO standards ensures an integrated approach to medical device safety, quality, and compatibility across various extracorporeal therapies.
Key SEO keywords: ISO 8637-3, plasmafilter standards, blood purification devices, plasmapheresis equipment, medical device sterility, single-use plasmafilters, medical device safety standards, ISO plasmafilter requirements, blood filter ISO standard.
Relations
- Effective Date
- 21-Sep-2024
Buy Documents
ISO/FDIS 8637-3 - Extracorporeal systems for blood purification — Part 3: Plasmafilters
REDLINE ISO/FDIS 8637-3 - Extracorporeal systems for blood purification — Part 3: Plasmafilters
Get Certified
Connect with accredited certification bodies for this standard

BSI Group
BSI (British Standards Institution) is the business standards company that helps organizations make excellence a habit.

TÜV Rheinland
TÜV Rheinland is a leading international provider of technical services.
TÜV SÜD
TÜV SÜD is a trusted partner of choice for safety, security and sustainability solutions.
Sponsored listings
Frequently Asked Questions
ISO/FDIS 8637-3 is a draft published by the International Organization for Standardization (ISO). Its full title is "Extracorporeal systems for blood purification — Part 3: Plasmafilters". This standard covers: This document specifies requirements and test methods for plasmafilters, which are devices intended to separate plasma from blood in therapeutic plasmapheresis therapy. This document specifies the requirements for sterile, single-use plasmafilters, intended for use on humans, hereinafter collectively referred to as “the device”, for use in humans. This document does not apply to; — extracorporeal blood circuits; — haemodialysers, haemodiafilters, haemofilters and haemoconcentrators; — haemoperfusion devices; — vascular access devices; — blood pumps; — systems or equipment intended to perform plasma separation. NOTE 1 Requirements for the extracorporeal blood circuit are specified in ISO 8637-2. NOTE 2 Requirements for haemodialysers, haemodiafilters, haemofilters and haemoconcentrators are specified in ISO 8637-1.
This document specifies requirements and test methods for plasmafilters, which are devices intended to separate plasma from blood in therapeutic plasmapheresis therapy. This document specifies the requirements for sterile, single-use plasmafilters, intended for use on humans, hereinafter collectively referred to as “the device”, for use in humans. This document does not apply to; — extracorporeal blood circuits; — haemodialysers, haemodiafilters, haemofilters and haemoconcentrators; — haemoperfusion devices; — vascular access devices; — blood pumps; — systems or equipment intended to perform plasma separation. NOTE 1 Requirements for the extracorporeal blood circuit are specified in ISO 8637-2. NOTE 2 Requirements for haemodialysers, haemodiafilters, haemofilters and haemoconcentrators are specified in ISO 8637-1.
ISO/FDIS 8637-3 is classified under the following ICS (International Classification for Standards) categories: 11.040.20 - Transfusion, infusion and injection equipment. The ICS classification helps identify the subject area and facilitates finding related standards.
ISO/FDIS 8637-3 has the following relationships with other standards: It is inter standard links to ISO 8637-3:2024. Understanding these relationships helps ensure you are using the most current and applicable version of the standard.
ISO/FDIS 8637-3 is available in PDF format for immediate download after purchase. The document can be added to your cart and obtained through the secure checkout process. Digital delivery ensures instant access to the complete standard document.
Standards Content (Sample)
FINAL DRAFT
International
Standard
ISO/TC 150/SC 2
Extracorporeal systems for blood
Secretariat: ANSI
purification —
Voting begins on:
2026-09-21
Part 3:
Plasmafilters
Voting terminates on:
2026-11-16
Systèmes extracorporels pour la purification du sang —
Partie 3: Filtres pour plasma
RECIPIENTS OF THIS DRAFT ARE INVITED TO SUBMIT,
WITH THEIR COMMENTS, NOTIFICATION OF ANY
RELEVANT PATENT RIGHTS OF WHICH THEY ARE AWARE
AND TO PROVIDE SUPPOR TING DOCUMENTATION.
IN ADDITION TO THEIR EVALUATION AS
BEING ACCEPTABLE FOR INDUSTRIAL, TECHNO-
LOGICAL, COMMERCIAL AND USER PURPOSES, DRAFT
INTERNATIONAL STANDARDS MAY ON OCCASION HAVE
TO BE CONSIDERED IN THE LIGHT OF THEIR POTENTIAL
TO BECOME STAN DARDS TO WHICH REFERENCE MAY BE
MADE IN NATIONAL REGULATIONS.
Reference number
FINAL DRAFT
International
Standard
ISO/TC 150/SC 2
Extracorporeal systems for blood
Secretariat: ANSI
purification —
Voting begins on:
Part 3:
Plasmafilters
Voting terminates on:
Systèmes extracorporels pour la purification du sang —
Partie 3: Filtres pour plasma
RECIPIENTS OF THIS DRAFT ARE INVITED TO SUBMIT,
WITH THEIR COMMENTS, NOTIFICATION OF ANY
RELEVANT PATENT RIGHTS OF WHICH THEY ARE AWARE
AND TO PROVIDE SUPPOR TING DOCUMENTATION.
© ISO 2026
IN ADDITION TO THEIR EVALUATION AS
All rights reserved. Unless otherwise specified, or required in the context of its implementation, no part of this publication may
BEING ACCEPTABLE FOR INDUSTRIAL, TECHNO-
LOGICAL, COMMERCIAL AND USER PURPOSES, DRAFT
be reproduced or utilized otherwise in any form or by any means, electronic or mechanical, including photocopying, or posting on
INTERNATIONAL STANDARDS MAY ON OCCASION HAVE
the internet or an intranet, without prior written permission. Permission can be requested from either ISO at the address below
TO BE CONSIDERED IN THE LIGHT OF THEIR POTENTIAL
or ISO’s member body in the country of the requester.
TO BECOME STAN DARDS TO WHICH REFERENCE MAY BE
MADE IN NATIONAL REGULATIONS.
ISO copyright office
CP 401 • Ch. de Blandonnet 8
CH-1214 Vernier, Geneva
Phone: +41 22 749 01 11
Email: copyright@iso.org
Website: www.iso.org
Published in Switzerland Reference number
ii
ISO/FDIS 8637-3:2026(en)
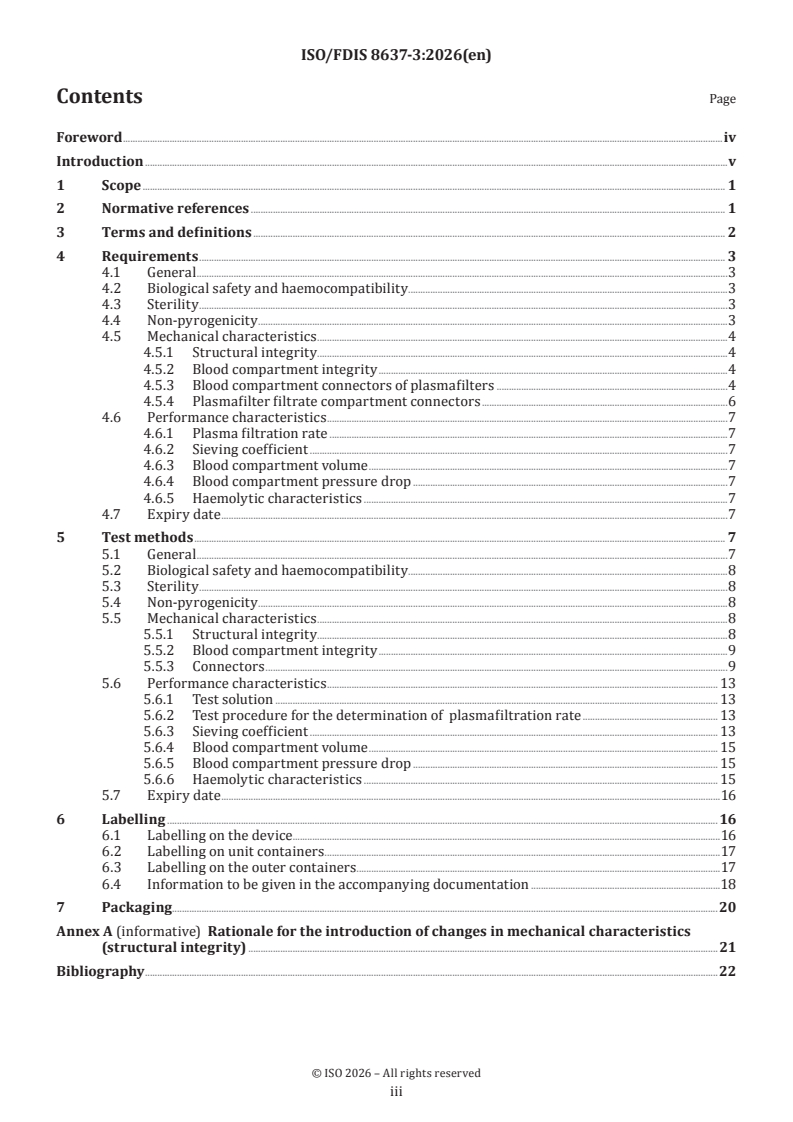
Contents Page
Foreword .iv
Introduction .v
1 Scope . 1
2 Normative references . 1
3 Terms and definitions . 2
4 Requirements . 3
4.1 General .3
4.2 Biological safety and haemocompatibility .3
4.3 Sterility .3
4.4 Non-pyrogenicity .3
4.5 Mechanical characteristics .4
4.5.1 Structural integrity.4
4.5.2 Blood compartment integrity .4
4.5.3 Blood compartment connectors of plasmafilters .4
4.5.4 Plasmafilter filtrate compartment connectors .6
4.6 Performance characteristics .7
4.6.1 Plasma filtration rate .7
4.6.2 Sieving coefficient .7
4.6.3 Blood compartment volume .7
4.6.4 Blood compartment pressure drop .7
4.6.5 Haemolytic characteristics .7
4.7 Expiry date .7
5 Test methods . 7
5.1 General .7
5.2 Biological safety and haemocompatibility .8
5.3 Sterility .8
5.4 Non-pyrogenicity .8
5.5 Mechanical characteristics .8
5.5.1 Structural integrity.8
5.5.2 Blood compartment integrity .9
5.5.3 Connectors .9
5.6 Performance characteristics . 13
5.6.1 Test solution . 13
5.6.2 Test procedure for the determination of plasmafiltration rate . 13
5.6.3 Sieving coefficient . 13
5.6.4 Blood compartment volume . 15
5.6.5 Blood compartment pressure drop . 15
5.6.6 Haemolytic characteristics . 15
5.7 Expiry date .16
6 Labelling .16
6.1 Labelling on the device.16
6.2 Labelling on unit containers .17
6.3 Labelling on the outer containers .17
6.4 Information to be given in the accompanying documentation .18
7 Packaging.20
Annex A (informative) Rationale for the introduction of changes in mechanical characteristics
(structural integrity) .21
Bibliography .22
iii
ISO/FDIS 8637-3:2026(en)
Foreword
ISO (the International Organization for Standardization) is a worldwide federation of national standards
bodies (ISO member bodies). The work of preparing International Standards is normally carried out through
ISO technical committees. Each member body interested in a subject for which a technical committee
has been established has the right to be represented on that committee. International organizations,
governmental and non-governmental, in liaison with ISO, also take part in the work. ISO collaborates closely
with the International Electrotechnical Commission (IEC) on all matters of electrotechnical standardization.
The procedures used to develop this document and those intended for its further maintenance are described
in the ISO/IEC Directives, Part 1. In particular, the different approval criteria needed for the different types
of ISO document should be noted. This document was drafted in accordance with the editorial rules of the
ISO/IEC Directives, Part 2 (see www.iso.org/directives).
ISO draws attention to the possibility that the implementation of this document may involve the use of (a)
patent(s). ISO takes no position concerning the evidence, validity or applicability of any claimed patent
rights in respect thereof. As of the date of publication of this document, ISO had not received notice of (a)
patent(s) which may be required to implement this document. However, implementers are cautioned that
this may not represent the latest information, which may be obtained from the patent database available at
www.iso.org/patents. ISO shall not be held responsible for identifying any or all such patent rights.
Any trade name used in this document is information given for the convenience of users and does not
constitute an endorsement.
For an explanation of the voluntary nature of standards, the meaning of ISO specific terms and expressions
related to conformity assessment, as well as information about ISO's adherence to the World Trade
Organization (WTO) principles in the Technical Barriers to Trade (TBT), see www.iso.org/iso/foreword.html.
This document was prepared by Technical committee ISO/TC 150, Implants for surgery, Subcommittee SC 2,
Cardiovascular implants and extracorporeal systems.
This third edition cancels and replaces the second edition (ISO 8637-3:2024), which has been technically
revised.
The main changes are as follows:
[1]
— addition of a link to ISO 14971 for risk management procedures;
— alignment of terms and definitions with those defined in other parts of the ISO 8637 series;
— revision of the procedure to demonstrate the properties required by the standard for the ss specification
of the expiration date;
— introduction of an Annex A to explain the rationale for the changes relating to mechanical strength in the
context of regulatory requirements.
A list of all the parts in the ISO 8637 series can be found on the ISO website.
Any feedback or questions on this document should be directed to the user’s national standards body. A
complete listing of these bodies can be found at www.iso.org/members.html.
iv
ISO/FDIS 8637-3:2026(en)
Introduction
This document is concerned with filters intended to perform plasma filtration in humans. If such a filter
is used with an extracorporeal circuit, the dimensions of the blood compartment connectors and filtrate
compartment connectors have been specified to ensure compatibility of the device with the extracorporeal
[2]
blood circuit specified in ISO 8637-2 . The design and dimensions have been selected to minimize the risk
of leakage of blood and the ingress of air.
There is no intention to specify, or to set limits on, the performance characteristics of the devices because
such restrictions are unnecessary for the qualified user and would limit the alternatives available when
choosing a device for a specific application.
All materials used in the construction of the device meet the necessary mechanical strength requirements as
well as those relating to biocompatibility, thrombogenicity, and pyrogenicity after sterilization. Therefore,
this document only requires that materials used have been tested, and that the testing methods and the test
results are made available upon request.
v
FINAL DRAFT International Standard ISO/FDIS 8637-3:2026(en)
Extracorporeal systems for blood purification —
Part 3:
Plasmafilters
1 Scope
This document specifies requirements and test methods for plasmafilters, which are devices intended
to separate plasma from blood in therapeutic plasmapheresis therapy. This document specifies the
requirements for sterile, single-use plasmafilters, intended for use on humans, hereinafter collectively
referred to as “the device”, for use in humans.
This document does not apply to
— extracorporeal blood circuits,
— haemodialysers, haemodiafilters, haemofilters and haemoconcentrators,
— haemoperfusion devices,
— vascular access devices,
— blood pumps, and
— systems or equipment intended to perform plasma separation.
[2]
NOTE 1 Requirements for the extracorporeal blood circuit are specified in ISO 8637-2 .
NOTE 2 Requirements for haemodialysers, haemodiafilters, haemofilters and haemoconcentrators are specified in
[3]
ISO 8637-1 .
2 Normative references
The following documents are referred to in the text in such a way that some or all of their content constitutes
requirements of this document. For dated references, only the edition cited applies. For undated references,
the latest edition of the referenced document (including any amendments) applies.
ISO/FDIS 8637-1, Extracorporeal systems for blood purification — Part 1: Haemodialysers, haemodiafilters,
haemofilters and haemoconcentrators
ISO 10993-1, Biological evaluation of medical devices — Part 1: Requirements and general principles for the
evaluation of biological safety within a risk management process
ISO 10993-4, Biological evaluation of medical devices — Part 4: Selection of tests for interactions with blood
ISO 10993-7, Biological evaluation of medical devices — Part 7: Ethylene oxide sterilization residuals
ISO 10993-11, Biological evaluation of medical devices — Part 11: Tests for systemic toxicity
ISO 11607-1, Packaging for terminally sterilized medical devices — Part 1: Requirements for materials, sterile
barrier systems and packaging systems
ISO 11607-2, Packaging for terminally sterilized medical devices — Part 2: Validation requirements for forming,
sealing and assembly processes
ISO/FDIS 8637-3:2026(en)
ISO 11737-2, Sterilization of health care products — Microbiological methods — Part 2: Tests of sterility
performed in the definition, validation and maintenance of a sterilization process
ISO 20417, Medical devices — Information to be supplied by the manufacturer
ISO 80369-7:2021, Small-bore connectors for liquids and gases in healthcare applications — Part 7: Connectors
for intravascular or hypodermic applications
ISO 80369-7, Small-bore connectors for liquids and gases in healthcare applications — Part 7: Connectors for
intravascular or hypodermic applications
ISO 80369-20, Small-bore connectors for liquids and gases in healthcare applications — Part 20: Common test
methods
3 Terms and definitions
For the purposes of this document, the following terms and definitions apply.
ISO and IEC maintain terminology databases for use in standardization at the following addresses:
— ISO Online browsing platform: available at https:// www .iso .org/ obp
— IEC Electropedia: available at https:// www .electropedia .org/
3.1
blood compartment
part of the plasmafilter (3.7) through which blood is intended to pass
3.2
blood compartment volume
volume which is needed to fill the blood compartment (3.1)
Note 1 to entry: For hollow fibre devices, the blood compartment volume (3.2) includes the volume of the hollow fibres
plus the headers.
3.3
plasma filtrate compartment
part of the plasmafilter (3.7) through which filtrate flows
3.4
plasma filtration rate
rate at which plasma is removed from the blood compartment (3.1) across the semipermeable membrane
into the plasma filtrate compartment (3.3) of a plasmafilter (3.7)
3.5
labelling
written, printed, graphic or electronic matter that is affixed to the plasmafilter (3.7) or any of its containers
or wrappers, or that accompanies a plasmafilter (3.7), and which is related to the identification, technical
description and use of that device, excluding shipping documents
3.6
plasma separation
plasmapheresis
plasma filtration
separation of a portion of the whole plasma from formed elements of blood by means of a semipermeable
membrane
Note 1 to entry: Plasma separation can also be accomplished through the use of centrifugation; however, this method
is not covered by this document.
ISO/FDIS 8637-3:2026(en)
3.7
plasmafilter
plasma separator
device intended to perform membrane plasmapheresis (3.6)
3.8
sieving coefficient
ratio of a solute concentration in the filtrate to the simultaneous concentration of the same solute on the
feed side of the plasmafilter
3.9
transmembrane pressure
TMP
p
TM
mean pressure exerted across the semipermeable membrane contained in the plasmafilter (3.7)
Note 1 to entry: The transmembrane pressure is given by Formula (1):
(1)
where
p is the pressure at the blood compartment inlet;
BI
p is the pressure at the blood compartment outlet;
BO
p is the pressure at the filtrate compartment outlet.
F
4 Requirements
4.1 General
This clause provides the requirements for a device evaluation of which shall conform to a structured
[1]
evaluation plan within a risk management process in accordance with ISO 14971 .
4.2 Biological safety and haemocompatibility
Parts of the plasmafilter that are intended to come into direct or indirect contact with blood shall be
evaluated for freedom from biological hazards in accordance with 5.2.
Attention is drawn to the importance of determining whether national regulations or national standards
governing toxicology and biocompatibility testing exist in the country in which the device is produced and, if
applicable, in the countries in which the device is to be marketed.
4.3 Sterility
The blood and filtrate pathways of the plasmafilter shall be sterile. The state of sterility of the device shall
conform with the manufacturer's statement [see 6.2, list item h)].
Conformity shall be verified in accordance with 5.3.
4.4 Non-pyrogenicity
The blood and filtrate pathways of the plasmafilter shall be non-pyrogenic and the state of pyrogenicity shall
comply with the manufacturer's statement [see 6.2, list item h)]. Conformity shall be verified in accordance
with 5.4.
ISO/FDIS 8637-3:2026(en)
4.5 Mechanical characteristics
4.5.1 Structural integrity
The device external casing shall be capable of withstanding the maximum positive pressure above
atmospheric pressure and the maximum negative sub-atmospheric pressure that can occur in the
plasma filtration system in accordance with the outputs of the manufacturer’s risk management process.
Considerations include, but are not limited to, the type of application, duration of treatment, pressures and
temperatures encountered, and foreseeable misuse. For basic safety, single failure shall also be considered
in the assessment. (See Annex A for further guidance.) Alternatively the maximum pressure recommended
by the manufacturer can be used to verify structural integrity.
Conformity shall be verified in accordance with 5.5.1.2 and 5.5.1.3.
4.5.2 Blood compartment integrity
When exposing the blood compartment of the device to a validated test procedure performed at a maximum
pressure derived from the manufacturer’s risk management process, the blood compartment shall not leak.
Test times and pressures can be determined based on the results of the product-specific risk management.
Considerations include, but are not limited to, the type of application, duration of treatment, pressures and
temperatures encountered, and foreseeable misuse. For basic safety, single failure shall also be considered
in the assessment. Alternatively the maximum pressure recommended by the manufacturer to verify
structural integrity can be used.
Conformity with this requirement shall be verified in accordance with 5.5.2.
4.5.3 Blood compartment connectors of plasmafilters
4.5.3.1 General
All connectors that connect the plasmafilter to the extracorporeal blood circuit shall provide a safe
connection. To ensure a safe connection, leakage of air from the outside or loss of blood to the environment
shall be avoided. The minimum separation force, minimum separation torque and maximum connection
torque shall be defined in accordance with the outputs of the manufacturer’s risk management process.
Boundary parameters used in tests such as torques, connection forces and disconnection forces, holding
times, and ambient temperatures shall be considered and defined as part of the manufacturer's assessment
on the use of the product. The selected forces and torques used in tests shall be representative of the
typical physical conditions of users. If necessary, occupational health and safety guidelines for maximum
permissible torques and forces should be taken into account.
4.5.3.2 Dimensional requirements
Except where the plasmafilter and the extracorporeal blood circuit are designed as an integral system, the
dimensions of the blood compartment connectors shall be as given in Figure 1 and Table 1.
Conformity with this requirement shall be verified in accordance with 5.5.3.2.
ISO/FDIS 8637-3:2026(en)
Figure 1 — Cone blood inlet and outlet blood compartment connectors of the plasmafilter
ISO/FDIS 8637-3:2026(en)
Table 1 — Blood compartment connector dimensions
a b c d
E F G H J K P α β γ
mm mm mm mm mm mm mm ° °
Minimum 10,8 0,85 5,97 — —
10 or 9 or 13 or
Nominal 8 11,0 1,1 6,0 15 15 6:100
more more more
Maximum 11,3 1,35 6,03 — —
Key
E length of tapered region
F length of tapered region
G thread pitch
H root diameter
J crest diameter
K thread crest width
P cone diameter
α angle of thread
β angle of thread
γ dimension taper rate
a
Double thread pitch.
b
Altered upper tolerance to accommodate different components and materials.
c
Revised dimension and tolerances based on existing manufacturing practice.
d
Cone's plane of reference: square A. Dimension measured as a projection on the front face. See Figure 1 (Z).
4.5.4 Plasmafilter filtrate compartment connectors
4.5.4.1 General
All connectors that connect to the plasmafilter filtrate compartment shall provide a safe and leak free
connection. Ingress of air from the outside or the leakage of filtrate to the environment shall be avoided. The
selected forces required to make the connection shall be representative of the typical physical conditions
of users. If necessary, occupational health and safety guidelines for maximum permissible forces should be
taken into account.
4.5.4.2 Dimensional requirements
Except when the plasmafilter and its extracorporeal circuit is designed as an integral system, the plasma
filtrate compartment connector shall be as follows:
a) a connector design in accordance with ISO/FDIS 8637-1:—, Figure 2 and Table 2;
b) a Luer lock connector design in accordance with ISO 80369-7:2021, Figures B.1 and B.3;
c) a non-locking connection for direct attachment of the tubing.
If non-locking connectors are used, they shall not separate under an axial force of 25 N applied for 15 s.
Filtrate connector functional requirements, such as acceptable leakage rate, minimum separation
force, minimum separation and maximum connection torque shall be defined in accordance with the
manufacturer’s risk management process. The boundary parameters used in tests such as the torques, the
connection and disconnection forces as well as holding times and ambient temperatures shall be considered
and defined as part of the manufacturer's risk assessment on the use of the product.
Functional testing shall be performed subject to the manufacturer's risk assessment.
Conformity with these requirements shall be verified in accordance with 5.5.3.3.2.
ISO/FDIS 8637-3:2026(en)
4.6 Performance characteristics
4.6.1 Plasma filtration rate
The plasma filtration rate shall be determined in accordance with 5.6.2. The blood flow rate shall cover the
manufacturers specified range for the plasmafilter (see 6.4).
4.6.2 Sieving coefficient
The sieving coefficients for albumin, immunoglobulin G (IgG), immunoglobulin M (IgM), apolipoprotein B
(apoB) or low density lipoprotein (LDL), or other equivalent indicators shall be determined in accordance
with 5.6.3.
4.6.3 Blood compartment volume
The volume of the blood compartment shall be determined in accordance with 5.6.4.
If the blood compartment volume is stable or constant over the clinical range of pressures, a single
measurement is sufficient. If the blood compartment volume varies with pressure, the blood compartment
volume over the clinical range of pressures shall be established.
4.6.4 Blood compartment pressure drop
The pressure drop of the blood compartment shall be determined in accordance with 5.6.5.
4.6.5 Haemolytic characteristics
The haemolytic characteristics shall be determined in accordance with 5.6.6.
4.7 Expiry date
The biological safety, sterility, performance data and mechanical integrity of the device shall be proven after
storage for a period corresponding to the expiry date.
Conformity shall be verified in accordance with 5.7.
5 Test methods
5.1 General
The requirements specified in 4.6 shall be determined prior to marketing a new type of plasmafilter and
shall be re-evaluated after changes in the device that can alter its performance.
For the tests, the device sample size shall be risk-based and shall be capable of demonstrating that the test
results meet the full range of specifications of the manufacturer with statistical confidence.
Configuration of the device samples used for the tests shall be representative of the final production
configuration, including sterilization.
Measurements shall be made in vitro at (37 ± 1) °C. When the relationship between variables is nonlinear,
sufficient determinations shall be made to permit interpolation between the data points. The techniques of
measurement given in this document are reference tests. Other test methods may be used, provided they
have been validated and shown to be precise and reproducible.
The test systems shown do not indicate all the necessary details of practicable test appara
...
ISO/TC 150/SC 2
Secretariat: ANSI
Date: 2026-07-01xx
Extracorporeal systems for blood purification —
Part 3:
Plasmafilters
Systèmes extracorporels pour la purification du sang —
Partie 3: Filtres pour plasma
FDIS stage
All rights reserved. Unless otherwise specified, or required in the context of its implementation, no part of this publication
may be reproduced or utilized otherwise in any form or by any means, electronic or mechanical, including photocopying,
or posting on the internet or an intranet, without prior written permission. Permission can be requested from either ISO
at the address below or ISO’s member body in the country of the requester.
ISO copyright office
CP 401 • Ch. de Blandonnet 8
CH-1214 Vernier, Geneva
Phone: + 41 22 749 01 11
E-mail: copyright@iso.org
Website: www.iso.org
Published in Switzerland
ii
Contents
Foreword . iv
Introduction . v
1 Scope . 1
2 Normative references . 1
3 Terms and definitions . 2
4 Requirements . 3
4.1 General . 3
4.2 Biological safety and haemocompatibility . 3
4.3 Sterility . 4
4.4 Non-pyrogenicity . 4
4.5 Mechanical characteristics . 4
4.6 Performance characteristics . 6
4.7 Expiry date . 7
5 Test methods . 7
5.1 General . 7
5.2 Biological safety and haemocompatibility . 8
5.3 Sterility . 8
5.4 Non-pyrogenicity . 8
5.5 Mechanical characteristics . 8
5.6 Performance characteristics . 13
5.7 Expiry date . 16
6 Labelling . 16
6.1 Labelling on the device . 16
6.2 Labelling on unit containers . 17
6.3 Labelling on the outer containers . 17
6.4 Information to be given in the accompanying documentation . 18
7 Packaging . 20
Annex A (informative) Rationale for the introduction of changes in mechanical characteristics
(structural integrity) . 21
Bibliography . 22
iii
Foreword
ISO (the International Organization for Standardization) is a worldwide federation of national standards
bodies (ISO member bodies). The work of preparing International Standards is normally carried out through
ISO technical committees. Each member body interested in a subject for which a technical committee has been
established has the right to be represented on that committee. International organizations, governmental and
non-governmental, in liaison with ISO, also take part in the work. ISO collaborates closely with the
International Electrotechnical Commission (IEC) on all matters of electrotechnical standardization.
The procedures used to develop this document and those intended for its further maintenance are described
in the ISO/IEC Directives, Part 1. In particular, the different approval criteria needed for the different types of
ISO document should be noted. This document was drafted in accordance with the editorial rules of the
ISO/IEC Directives, Part 2 (see www.iso.org/directives).
ISO draws attention to the possibility that the implementation of this document may involve the use of (a)
patent(s). ISO takes no position concerning the evidence, validity or applicability of any claimed patent rights
in respect thereof. As of the date of publication of this document, ISO had not received notice of (a) patent(s)
which may be required to implement this document. However, implementers are cautioned that this may not
represent the latest information, which may be obtained from the patent database available at
www.iso.org/patents. ISO shall not be held responsible for identifying any or all such patent rights.
Any trade name used in this document is information given for the convenience of users and does not
constitute an endorsement.
For an explanation of the voluntary nature of standards, the meaning of ISO specific terms and expressions
related to conformity assessment, as well as information about ISO's adherence to the World Trade
Organization (WTO) principles in the Technical Barriers to Trade (TBT), see www.iso.org/iso/foreword.html.
This document was prepared by Technical committee ISO/TC 150, Implants for surgery, Subcommittee SC 2,
Cardiovascular implants and extracorporeal systems.
This third edition cancels and replaces the second edition (ISO 8637-3:2024), which has been technically
revised.
The main changes are as follows:
[1]
— addition of a link to ISO 14971 ISO 14971 for risk management procedures;
— alignment of terms and definitions with those defined in other parts of the ISO 8637 series;
— revision of the procedure to demonstrate the properties required by the standard for the ss specification
of the expiration date;
— introduction of an Annex A to explain the rationale for the changes relating to mechanical strength in the
context of regulatory requirements.
A list of all the parts in the ISO 8637 series can be found on the ISO website.
Any feedback or questions on this document should be directed to the user’s national standards body. A
complete listing of these bodies can be found at www.iso.org/members.html.
iv
Introduction
This document is concerned with filters intended to perform plasma filtration () in humans. If such a filter is
used with an extracorporeal circuit, the dimensions of the blood compartment connectors and filtrate
compartment connectors have been specified to ensure compatibility of the device with the extracorporeal
[2]
blood circuit specified in ISO 8637-2 . The design and dimensions have been selected to minimize the risk
of leakage of blood and the ingress of air.
There is no intention to specify, or to set limits on, the performance characteristics of the devices because such
restrictions are unnecessary for the qualified user and would limit the alternatives available when choosing a
device for a specific application.
All materials used in the construction of the device meet the necessary mechanical strength requirements as
well as those relating to biocompatibility, thrombogenicity, and pyrogenicity after sterilization. Therefore, this
document only requires that materials used have been tested, and that the testing methods and the test results
are made available upon request.
v
Extracorporeal systems for blood purification —
Part 3:
Plasmafilters
1 Scope
This document specifies requirements and test methods for plasmafilters, which are devices intended to
separate plasma from blood in therapeutic plasmapheresis therapy. This document specifies the requirements
for sterile, single-use plasmafilters, intended for use on humans, hereinafter collectively referred to as “the
device”, for use in humans.
This document does not apply to;
— extracorporeal blood circuits;,
— haemodialysers, haemodiafilters, haemofilters and haemoconcentrators;,
— haemoperfusion devices;,
— vascular access devices;,
— blood pumps;, and
— systems or equipment intended to perform plasma separation.
[2]
NOTE 1 Requirements for the extracorporeal blood circuit are specified in ISO 8637-2 .
NOTE 2 Requirements for haemodialysers, haemodiafilters, haemofilters and haemoconcentrators are specified in
[3]
ISO 8637-1 .
2 Normative references
The following documents are referred to in the text in such a way that some or all of their content constitutes
requirements of this document. For dated references, only the edition cited applies. For undated references,
the latest edition of the referenced document (including any amendments) applies.
ISO 7000, Graphical symbols for use on equipment — Registered symbols
ISO/FDIS 8637-1, Extracorporeal systems for blood purification — Part 1: Haemodialysers, haemodiafilters,
haemofilters and haemoconcentrators
ISO 10993-1:2025, Biological evaluation of medical devices — Part 1: Requirements and general principles for
the evaluation of biological safety within a risk management process
ISO 10993-4, Biological evaluation of medical devices — Part 4: Selection of tests for interactions with blood
ISO 10993-7, Biological evaluation of medical devices — Part 7: Ethylene oxide sterilization residuals
ISO 10993-11, Biological evaluation of medical devices — Part 11: Tests for systemic toxicity
ISO 11607-1, Packaging for terminally sterilized medical devices — Part 1: Requirements for materials, sterile
barrier systems and packaging systems
ISO 11607-2, Packaging for terminally sterilized medical devices — Part 2: Validation requirements for forming,
sealing and assembly processes
ISO 11737-2:2019, Sterilization of health care products — Microbiological methods — Part 2: Tests of sterility
performed in the definition, validation and maintenance of a sterilization process
ISO 15223-1, Medical devices — Symbols to be used with information to be supplied by the manufacturer — Part
1: General requirements
ISO 20417, Medical devices — Information to be supplied by the manufacturer
ISO 80369-7:2021, Small-bore connectors for liquids and gases in healthcare applications — Part 7: Connectors
for intravascular or hypodermic applications
ISO 80369-7, Small-bore connectors for liquids and gases in healthcare applications — Part 7: Connectors for
intravascular or hypodermic applications
ISO 80369-20, Small-bore connectors for liquids and gases in healthcare applications — Part 20: Common test
methods
3 Terms and definitions
For the purposes of this document, the following terms and definitions apply.
ISO and IEC maintain terminology databases for use in standardization at the following addresses:
— ISO Online browsing platform: available at https://www.iso.org/obp
— IEC Electropedia: available at https://www.electropedia.org/
3.1
blood compartment, noun
part of the plasmafilter (3.7) through which blood is intended to pass
3.2
blood compartment volume, noun
volume which is needed to fill the blood compartment (3.1)
Note 1 to entry: For hollow fibre devices, the blood compartment volume (3.2) includes the volume of the hollow fibres
plus the headers.
3.3
plasma filtrate compartment, noun
part of the plasmafilter (3.7) through which filtrate flows
3.4
plasma filtration rate, noun
rate at which plasma is removed from the blood compartment (3.1) across the semipermeable membrane into
the plasma filtrate compartment (3.3) of a plasmafilter (3.7)
3.5
labelling, noun
written, printed, graphic or electronic matter that is affixed to the plasmafilter (3.7) or any of its containers or
wrappers, or that accompanies a plasmafilter (3.7), and which is related to the identification, technical
description and use of that device, excluding shipping documents
3.6
plasma separation, noun
plasmapheresis, noun
plasma filtration, noun
separation of a portion of the whole plasma from formed elements of blood by means of a semipermeable
membrane
Note 1 to entry: Plasma separation can also be accomplished through the use of centrifugation; however, this method is
not covered by this document.
3.7
plasmafilter, noun
plasma separator, noun
device intended to perform membrane plasmapheresis (3.6)
3.8
sieving coefficient, noun
ratio of a solute concentration in the filtrate to the simultaneous concentration of the same solute on the feed
side of the plasmafilter
3.9
transmembrane pressure, noun
TMP, noun
p , noun
TM
mean pressure exerted across the semipermeable membrane contained in the plasmafilter (3.7)
Note 1 to entry: The transmembrane pressure is given by Formula (1):
𝑝𝑝 +𝑝𝑝
BI BO
𝑝𝑝 = −𝑝𝑝 (1)
TM F
where
p is the pressure at the blood compartment inlet;
BI
p is the pressure at the blood compartment outlet;
BO
p is the pressure at the filtrate compartment outlet.
F
4 Requirements
4.1 General
This clause provides the requirements for a device evaluation of which shall conform to a structured
[1]
evaluation plan within a risk management process in accordance with ISO 14971 .
4.2 Biological safety and haemocompatibility
Parts of the plasmafilter that are intended to come into direct or indirect contact with blood shall be evaluated
for freedom from biological hazards, in accordance with 5.2.
Attention is drawn to the importance of determining whether national regulations or national standards
governing toxicology and biocompatibility testing exist in the country in which the device is produced and, if
applicable, in the countries in which the device is to be marketed.
4.3 Sterility
The blood and filtrate pathways of the plasmafilter shall be sterile. And theThe state of sterility of the device
shall conform with the manufacturer's statement [see 6.2, list item h).
Conformance.Conformity shall be verified in accordance with 5.3.
4.4 Non-pyrogenicity
The blood and filtrate pathways of the plasmafilter shall be non-pyrogenic and the state of pyrogenicity shall
comply with the manufacturer's statement [see 6.2, list item h). ComplianceConformity shall be verified in
accordance with 5.4.
4.5 Mechanical characteristics
4.5.1 Structural integrity
The device external casing shall be capable of withstanding the maximum positive pressure above
atmospheric pressure and the maximum negative sub-atmospheric pressure that can occur in the plasma
filtration system in accordance with the outputs of the manufacturer’s risk management process.
Considerations include, but are not limited to, the type of application, duration of treatment, pressures and
temperatures encountered, and foreseeable misuse. For basic safety, single failure shall also be considered in
the assessment. (See Annex A for further guidance.) Alternatively the maximum pressure recommended by
the manufacturer can be used to verify structural integrity.
Conformity shall be verified in accordance with 5.5.1.2 and 5.5.1.3.
4.5.2 Blood compartment integrity
When exposing the blood compartment of the device to a validated test procedure performed at a maximum
pressure derived from the manufacturer’s risk management process, the blood compartment shall not leak.
Test times and pressures can be determined based on the results of the product-specific risk management.
Considerations include, but are not limited to, the type of application, duration of treatment, pressures and
temperatures encountered, and foreseeable misuse. For basic safety, single failure mustshall also be
considered in the assessment. Alternatively the maximum pressure recommended by the manufacturer to
verify structural integrity can be used.
ComplianceConformity with this requirement shall be verified in accordance with 5.5.2.
4.5.3 Blood compartment connectors of plasmafilters
4.5.3.1 General
All connectors that connect the plasmafilter to the extracorporeal blood circuit shall provide a safe connection.
To ensure a safe connection, leakage of air from the outside or loss of blood to the environment shall be
avoided. The minimum separation force, minimum separation torque and maximum connection torque shall
be defined in accordance with the outputs of the manufacturer’s risk management process. Boundary
parameters used in tests such as torques, connection forces and disconnection forces, holding times, and
ambient temperatures, shall be considered and defined as part of the manufacturer's assessment on the use
of the product. The selected forces and torques used in tests shall be representative of the typical physical
conditions of users. If necessary, occupational health and safety guidelines for maximum permissible torques
and forces should be taken into account.
4.5.3.2 Dimensional requirements
Except where the plasmafilter and the extracorporeal blood circuit are designed as an integral system, the
dimensions of the blood compartment connectors shall be as given in Figure 1 and Table 1.
ComplianceConformity with this requirement shall be verified in accordance with 5.5.3.2.
Figure 1 — Cone blood inlet and outlet blood compartment connectors of the plasmafilter
Table 1 — Blood compartment connector dimensions
a b c d
E F G H J K P α β γ
mm mm mm mm mm mm mm ° °
Minimum 10,8 0,85 5,97 — —
10 or 13 or
Nominal 9 or more 8 11,0 1,1 6,0 15 15 6:100
more more
Maximum 11,3 1,35 6,03 — —
Key
E length of tapered region
F length of tapered region
G thread pitch
a b c d
E F G H J K P α β γ
mm mm mm mm mm mm mm ° °
H root diameter
J crest diameter
K thread crest width
P cone diameter
α angle of thread
β angle of thread
γ dimension taper rate
a
Double thread pitch.
b
Altered upper tolerance to accommodate different components and materials.
c
Revised dimension and tolerances based on existing manufacturing practice.
d
Cone's plane of reference: square A. Dimension measured as a projection on the front face. See Figure 1 (Z).
4.5.4 Plasmafilter filtrate compartment connectors
4.5.4.1 General
All connectors that connect to the plasmafilter filtrate compartment shall provide a safe and leak free
connection. Ingress of air from the outside or the leakage of filtrate to the environment shall be avoided. The
selected forces required to make the connection shall be representative of the typical physical conditions of
users. If necessary, occupational health and safety guidelines for maximum permissible forces should be taken
into account.
4.5.4.2 Dimensional requirements
Except when the plasmafilter and its extracorporeal circuit is designed as an integral system, the plasma
filtrate compartment connector shall be as follows:
a) a connector design in accordance with ISO/FDIS 8637-1:—, Figure 2 and Table 2; or
b) a Luer lock connector design in accordance with ISO 80369-7:2021, Figures B.1 and B.3; or
c) a non-locking connection for direct attachment of the tubing.
If non-locking connectors are used, they shall not separate under an axial force of 25 N applied for 15 s.
Filtrate connector functional requirements, such as acceptable leakage rate, minimum separation force,
minimum separation and maximum connection torque shall be defined in accordance with the manufacturer’s
risk management process. The boundary parameters used in tests such as the torques, the connection and
disconnection forces as well as holding times and ambient temperatures mustshall be considered and defined
as part of the manufacturer's risk assessment on the use of the product.
Functional testing shall be performed subject to the manufacturer's risk assessment.
ComplianceConformity with these requirements shall be verified in accordance with 5.5.3.3.2.
4.6 Performance characteristics
4.6.1 Plasma filtration rate
The plasma filtration rate shall be determined in accordance with 5.6.2. The blood flow rate shall cover the
manufacturers specified range for the plasmafilter (see 6.4).
4.6.2 Sieving coefficient
The sieving coefficients for albumin, immunoglobulin G (IgG), immunoglobulin M (IgM), apolipoprotein B
(apoB) or low density lipoprotein (LDL), or other equivalent indicators shall be determined in accordance
with 5.6.3.
4.6.3 Blood compartment volume
The volume of the blood compartment shall be determined in accordance with 5.6.4.
If the blood compartment volume is stable or constant over the clinical range of pressures, a single
measurement is sufficient. If the blood compartment volume varies with pressure, the blood compartment
volume over the clinical range of pressures shall be established.
4.6.4 Blood compartment pressure drop
The pressure drop of the blood compartment shall be determined in accordance with 5.6.5.
4.6.5 Haemolytic characteristics
The haemolytic characteristics shall be determined in accordance with 5.6.6.
4.7 Expiry date
The biological safety, sterility, performance data and mechanical integrity of the device shall be proven after
storage for a period corresponding to the expiry date.
ComplianceConformity shall be verified in accordance with 5.7.
5 Test methods
5.1 General
The requirements specified in 4.6 shall be determined prior to marketing a new type of plasmafilter and shall
be re-evaluated after changes in the device that can alter its performance.
For the tests, the device sample size shall be risk-based and shall be capable of demonstrating that the test
results meet the full range of specifications of the manufacturer with statistical confidence.
Configuration of the device samples used for the tests shall be representative of the final production
configuration, including sterilization.
Measurements shall be made in vitro at (37 ± 1) °C. When the relationship between variables is nonlinear,
sufficient determinations shall be made to permit interpolation between the data points. The techniques of
measurement given in this document are reference tests. Other test methods may be used, provided they have
been validated and shown to be precise and reproducible.
The test systems shown do not indicate all the necessary details of practicable test apparatus. The design and
construction of actual test systems shall also address factors contributing to measurement error, including,
but not limited to, pressure measurement errors due to static head effects and dynamic pressure drops,
parameter stabilization time, uncontrolled temperature variations, pH, degradation of test substances due to
heat, light and time, degassing of test fluids, trapped air, and system contamination by foreign material, algae
and bacteria.
NOTE This clause contains tests that are of a type-testing nature, which are carried out prior to marketing of a new
device or when changes are made to the device or its manufacturing processes. Other tests in this clause are of a quality
control nature, which are repeated on a regular basis according to quality management system requirements.
5.2 Biological safety and haemocompatibility
The biological safety of plasmafilter pathways that are intended to come into direct or indirect contact with
the patient's blood shall be evaluated on samples of each new type of device prior to its marketing, or after
any change in the materials of construction of that type of device, or after any change in the method of
sterilization. Testing shall be carried out in accordance with ISO 10993-1:2025, ISO 10993-4, ISO 10993-7 and
ISO 10993-11, as relevant.
5.3 Sterility
ComplianceConformity with 4.3 shall be verified by the inspection of the records to show that the device has
been exposed to a sterilization process that has been validated by ISO 11737-2:2019.
5.4 Non-pyrogenicity
ConformanceConformity with 4.4 shall be verified in accordance with ISO 10993-11.
NOTE ISO 10993-11ISO ISO 10993-11 does not specifically address requirements for endotoxin mediated
[5]
pyrogenicity test methods but makes reference to ANSI/AAMI ST72 ANSI/AAMI ST72.
5.5 Mechanical characteristics
5.5.1 Structural integrity
5.5.1.1 General
The requirements of 4.5.1 shall be verified by the following test methods together with reference to Annex A
for further guidance.
5.5.1.2 Structural integrity under a positive pressure
Completely fill the device with degassed water at (37 ± 1) °C. Seal all connectors except the connector to which
pressure is applied. Apply the maximum positive pressure that can occur in the plasma filtration system in
accordance with the outputs of the manufacturer’s risk management process or a positive air pressure
1,5 times of the manufacturer's recommended maximum pressure and seal the apparatus. After a period of
time derived from the manufacturer’s risk management process or at least 10 min, record the pressure and
visually examine the device for leaks.
Alternately, a constant air pressure (maximum positive pressure that can occur in the plasma filtration system
in accordance with the outputs of the manufacturer’s risk management process or 1,5 times of the
manufacturer’s recommended maximum pressure) can be applied and the device can be submerged in water
to test for air leakage.
Equivalent or superior tests may be used if available.
5.5.
...